Raman signatures of monoclinic distortion in (Ba1-xSrx)(3)CaNb2O9 complex perovskites
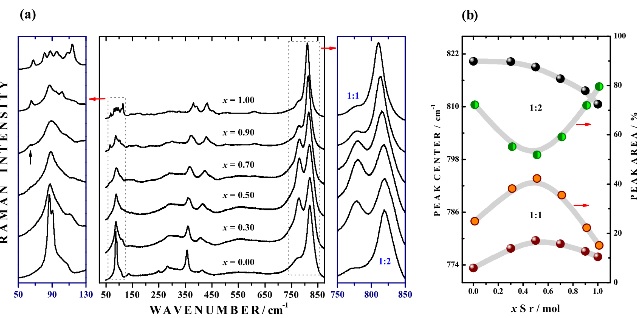
Abstract: Octahedral tilting is the most common distortion process observed in centrosymmetric perovskite compounds. Indeed, crucial physical properties of this oxide stem from the tilts of BO6 rigid octahedra. In microwave ceramics with perovskite-type structure, there is a close relation between the temperature coefficient of resonant frequency and the tilt system of the perovskite structure. However, in many cases, limited access facilities are needed to assign correctly the space group, including neutron scattering and transmission electron microscopy. Here, we combine the Raman scattering and group theory calculations to probe the structural distortion in the perovskite (Ba1-xSrx)(3)CaNb2O9 solid solution, which exhibits a structural phase transition at x0.7, from D-3d(3) trigonal to C-2h(3) monoclinic cell. Both phases are related by an octahedral tilting distortion (a(0)b(-)b(-) in Glazer notation). Low-temperature Raman spectra corroborate the group-theoretical predictions for Sr3CaNb2O9 compound because 36 modes detected at 25K agree well with the 42 (25A(g)circle plus 17B(g)) predicted ones. Copyright (c) 2017 John Wiley & Sons, Ltd.
Author(s): Rodrigues, J. E.; Bezerra, D. M.; Costa, R. C.; et al.
Journal of Raman Spectroscopy
Volume: 48 Issue: 9 Pages: 1243-1249 Published: 2017
DOI: http://dx.doi.org/10.1002/jrs.5195
PDF: Raman signatures of monoclinic distortion in (Ba1-xSrx)(3)CaNb2O9 complex perovskites